Revision Real-data Analysis R6: enhancer enrichment of FASH dynamic eQTLs
Ziang Zhang
2026-08-26
Last updated: 2026-08-26
Checks: 6 1
Knit directory: coderepo-local/
This reproducible R Markdown analysis was created with workflowr (version 1.7.2). The Checks tab describes the reproducibility checks that were applied when the results were created. The Past versions tab lists the development history.
The R Markdown file has unstaged changes. To know which version of
the R Markdown file created these results, you’ll want to first commit
it to the Git repo. If you’re still working on the analysis, you can
ignore this warning. When you’re finished, you can run
wflow_publish to commit the R Markdown file and build the
HTML.
Great job! The global environment was empty. Objects defined in the global environment can affect the analysis in your R Markdown file in unknown ways. For reproduciblity it’s best to always run the code in an empty environment.
The command set.seed(20251109) was run prior to running
the code in the R Markdown file. Setting a seed ensures that any results
that rely on randomness, e.g. subsampling or permutations, are
reproducible.
Great job! Recording the operating system, R version, and package versions is critical for reproducibility.
Nice! There were no cached chunks for this analysis, so you can be confident that you successfully produced the results during this run.
Great job! Using relative paths to the files within your workflowr project makes it easier to run your code on other machines.
Great! You are using Git for version control. Tracking code development and connecting the code version to the results is critical for reproducibility.
The results in this page were generated with repository version 55cacb9. See the Past versions tab to see a history of the changes made to the R Markdown and HTML files.
Note that you need to be careful to ensure that all relevant files for
the analysis have been committed to Git prior to generating the results
(you can use wflow_publish or
wflow_git_commit). workflowr only checks the R Markdown
file, but you know if there are other scripts or data files that it
depends on. Below is the status of the Git repository when the results
were generated:
Ignored files:
    Ignored:    .DS_Store
    Ignored:    .Rhistory
    Ignored:    .Rproj.user/
    Ignored:    Rplots.pdf
    Ignored:    analysis/.DS_Store
    Ignored:    analysis/.Rhistory
    Ignored:    code/.DS_Store
    Ignored:    code/.Rhistory
    Ignored:    code/revision_simulations/.DS_Store
    Ignored:    data/appendixB/
    Ignored:    data/dynamic_eQTL_real/
    Ignored:    data/revision_simulations/
    Ignored:    data/toy_example/
    Ignored:    output/dynamic_eQTL_real/
    Ignored:    output/pdf/
    Ignored:    output/playwright/
    Ignored:    output/revision_simulations/
    Ignored:    scripts/

Untracked files:
    Untracked:  analysis/revision_internal_fash_knockoff.rmd
    Untracked:  analysis/revision_internal_fash_temporal_class_enhancer_enrichment.rmd
    Untracked:  analysis/revision_internal_three_likelihood_comparison.rmd
    Untracked:  code/03_nonlindyn_lfsr.R
    Untracked:  code/03_nonlindyn_lfsr.sbatch
    Untracked:  code/revision_simulations/internal/fash_knockoff/reporting.R
    Untracked:  code/revision_simulations/internal/fash_knockoff/test_reporting.R
    Untracked:  code/revision_simulations/internal/fash_temporal_class_enhancer_enrichment/
    Untracked:  code/revision_simulations/internal/per_unit_expression_correlation/
    Untracked:  code/revision_simulations/r1_r2_fashr0143/source_snapshots/
    Untracked:  code/revision_simulations/r3_r4_fashr0143/21_r3_center_aligned_relative_location_paired_core.R
    Untracked:  code/revision_simulations/r3_r4_fashr0143/21_r3_center_aligned_relative_location_paired_fashr0143.R
    Untracked:  code/revision_simulations/r3_r4_fashr0143/21_r3_center_aligned_relative_location_paired_fashr0143.sbatch
    Untracked:  code/revision_simulations/r3_r4_fashr0143/source_snapshots/r3_center_aligned_relative_location_paired_driver.R
    Untracked:  code/revision_simulations/r3_r4_fashr0143/source_snapshots/r3_center_aligned_relative_location_paired_simulation_functions.R
    Untracked:  code/revision_simulations/r3_r4_fashr0143/test_r3_center_aligned_relative_location_paired.R
    Untracked:  code/revision_simulations/r4_correlated_errors/compute_full_model_residual_correlation.R
    Untracked:  code/revision_simulations/r4_correlated_errors/compute_pc_residual_expression_correlation.R

Unstaged changes:
    Modified:   analysis/dynamic_eQTL_real_full.rmd
    Modified:   analysis/nonlinear_dynamic_eQTL_real_full.rmd
    Modified:   analysis/revision_combined_spiky_genotype_simulation.rmd
    Modified:   analysis/revision_correlated_error_simulation.rmd
    Modified:   analysis/revision_enhancer_enrichment.rmd
    Modified:   analysis/revision_genotype_level_simulation.rmd
    Modified:   analysis/revision_internal_temporal_class_roadmap_enrichment.rmd
    Deleted:    code/02_dyn.R
    Modified:   code/02_dyn_lfsr.R
    Modified:   code/02_dyn_lfsr.sbatch
    Deleted:    code/03_nonlindyn.R
    Modified:   code/revision_simulations/README.md
    Modified:   code/revision_simulations/internal/fash_knockoff/plot_lfdr_histograms.R
    Modified:   code/revision_simulations/internal/fash_knockoff/plot_w_boxplot_by_discovery.R
    Modified:   code/revision_simulations/internal/fash_strober_enhancer_comparison/reporting.R
    Modified:   code/revision_simulations/internal/temporal_class_roadmap_enrichment/reporting.R
    Modified:   code/revision_simulations/internal/temporal_class_roadmap_enrichment/run_temporal_class_roadmap_enrichment.R
    Modified:   code/revision_simulations/internal/temporal_class_roadmap_enrichment/temporal_class_roadmap_helpers.R
    Modified:   code/revision_simulations/internal/temporal_class_roadmap_enrichment/test_temporal_class_roadmap_helpers.R
    Modified:   code/revision_simulations/r1_r2_fashr0143/reporting_overlay.R
    Modified:   code/revision_simulations/r4_correlated_errors/reporting.R
    Modified:   code/revision_simulations/r4_correlated_errors/test_unit_specific_residual_permutation_helpers.R
    Modified:   code/revision_simulations/r4_correlated_errors/unit_specific_residual_permutation_helpers.R

Staged changes:
    New:        analysis/revision_internal_temporal_class_roadmap_enrichment.rmd
    New:        code/revision_simulations/internal/temporal_class_roadmap_enrichment/reporting.R
    New:        code/revision_simulations/internal/temporal_class_roadmap_enrichment/run_temporal_class_roadmap_enrichment.R
    New:        code/revision_simulations/internal/temporal_class_roadmap_enrichment/temporal_class_roadmap_helpers.R
    New:        code/revision_simulations/internal/temporal_class_roadmap_enrichment/test_reporting.R
    New:        code/revision_simulations/internal/temporal_class_roadmap_enrichment/test_temporal_class_roadmap_helpers.R
Note that any generated files, e.g. HTML, png, CSS, etc., are not included in this status report because it is ok for generated content to have uncommitted changes.
These are the previous versions of the repository in which changes were
made to the R Markdown
(analysis/revision_enhancer_enrichment.rmd) and HTML
(docs/revision_enhancer_enrichment.html) files. If you’ve
configured a remote Git repository (see ?wflow_git_remote),
click on the hyperlinks in the table below to view the files as they
were in that past version.
| File | Version | Author | Date | Message |
|---|---|---|---|---|
| html | 83ed337 | Ziang Zhang | 2026-08-25 | Build site. |
| Rmd | 052df2a | Ziang Zhang | 2026-08-25 | Simplify R6 enhancer-enrichment page to three Roadmap annotations |
| Rmd | 09f63bc | Ziang Zhang | 2026-08-24 | Sync revision analyses and rendered pages |
| html | 09f63bc | Ziang Zhang | 2026-08-24 | Sync revision analyses and rendered pages |
| Rmd | c87957a | Ziang Zhang | 2026-08-10 | Add revision analyses: R1-R6 reviewer-facing pages and internal studies |
| html | c87957a | Ziang Zhang | 2026-08-10 | Add revision analyses: R1-R6 reviewer-facing pages and internal studies |
Question
Do the dynamic eQTLs that FASH discovers fall in functionally relevant enhancers? As an orthogonal biological check, we ask whether one lead variant per discovered gene overlaps established Roadmap enhancer annotations more often than the variants tested in this study do. We consider Roadmap epigenomes relevant to pluripotent, germ-layer, cardiac, and related vascular states, and use the enhancer definition adopted by Strober et al. (2019). The estimates below are descriptive and exploratory: they are not a fully adjusted functional enrichment analysis, and they are not evidence that any individual discovery is correct.
Two sections, both using one lead variant per discovered gene:
- Section 1 compares current FASH with Strober’s linear and quadratic/nonlinear analyses.
- Section 2 compares all FASH lead variants with the subset at rsIDs that neither Strober analysis identified.
Roadmap enhancer annotations
Following Strober et al. (2019), enhancer regions were defined using
the Roadmap 15-state ChromHMM enhancer annotations (Enh,
EnhG, EnhBiv, and BivFlnk)
(Roadmap Epigenomics Consortium et al. 2015). Using an established
annotation procedure avoids introducing an enhancer definition of our
own. Thirteen epigenomes are displayed, in four groups:
| Group | Roadmap ID | Roadmap tissue / cell type |
|---|---|---|
| iPSC | E018 | iPS-15b Cells |
| iPSC | E019 | iPS-18 Cells |
| iPSC | E020 | iPS-20b Cells |
| iPSC | E021 | iPS DF 6.9 Cells |
| iPSC | E022 | iPS DF 19.11 Cells |
| Germ layer | E011 | hESC Derived CD184+ Endoderm Cultured Cells |
| Germ layer | E012 | hESC Derived CD56+ Ectoderm Cultured Cells |
| Germ layer | E013 | hESC Derived CD56+ Mesoderm Cultured Cells |
| Cardiac | E083 | Fetal Heart |
| Cardiac | E095 | Left Ventricle |
| Cardiac | E104 | Right Atrium |
| Cardiac | E105 | Right Ventricle |
| Vascular | E065 | Aorta |
The iPSC and heart epigenome choices follow Strober et al. (2019); the three hESC-derived germ-layer epigenomes are included because cardiomyocytes arise through mesoderm. E065 aorta is a vascular rather than a cardiac tissue, so it is kept as a separate, related vascular annotation and is never counted among the cardiac epigenomes. Each tested variant is treated as a single base and intersected with the published Roadmap enhancer intervals, giving one indicator per variant per epigenome over all 745,867 tested variants.
Section 1: FASH versus Strober
plot_method_comparison()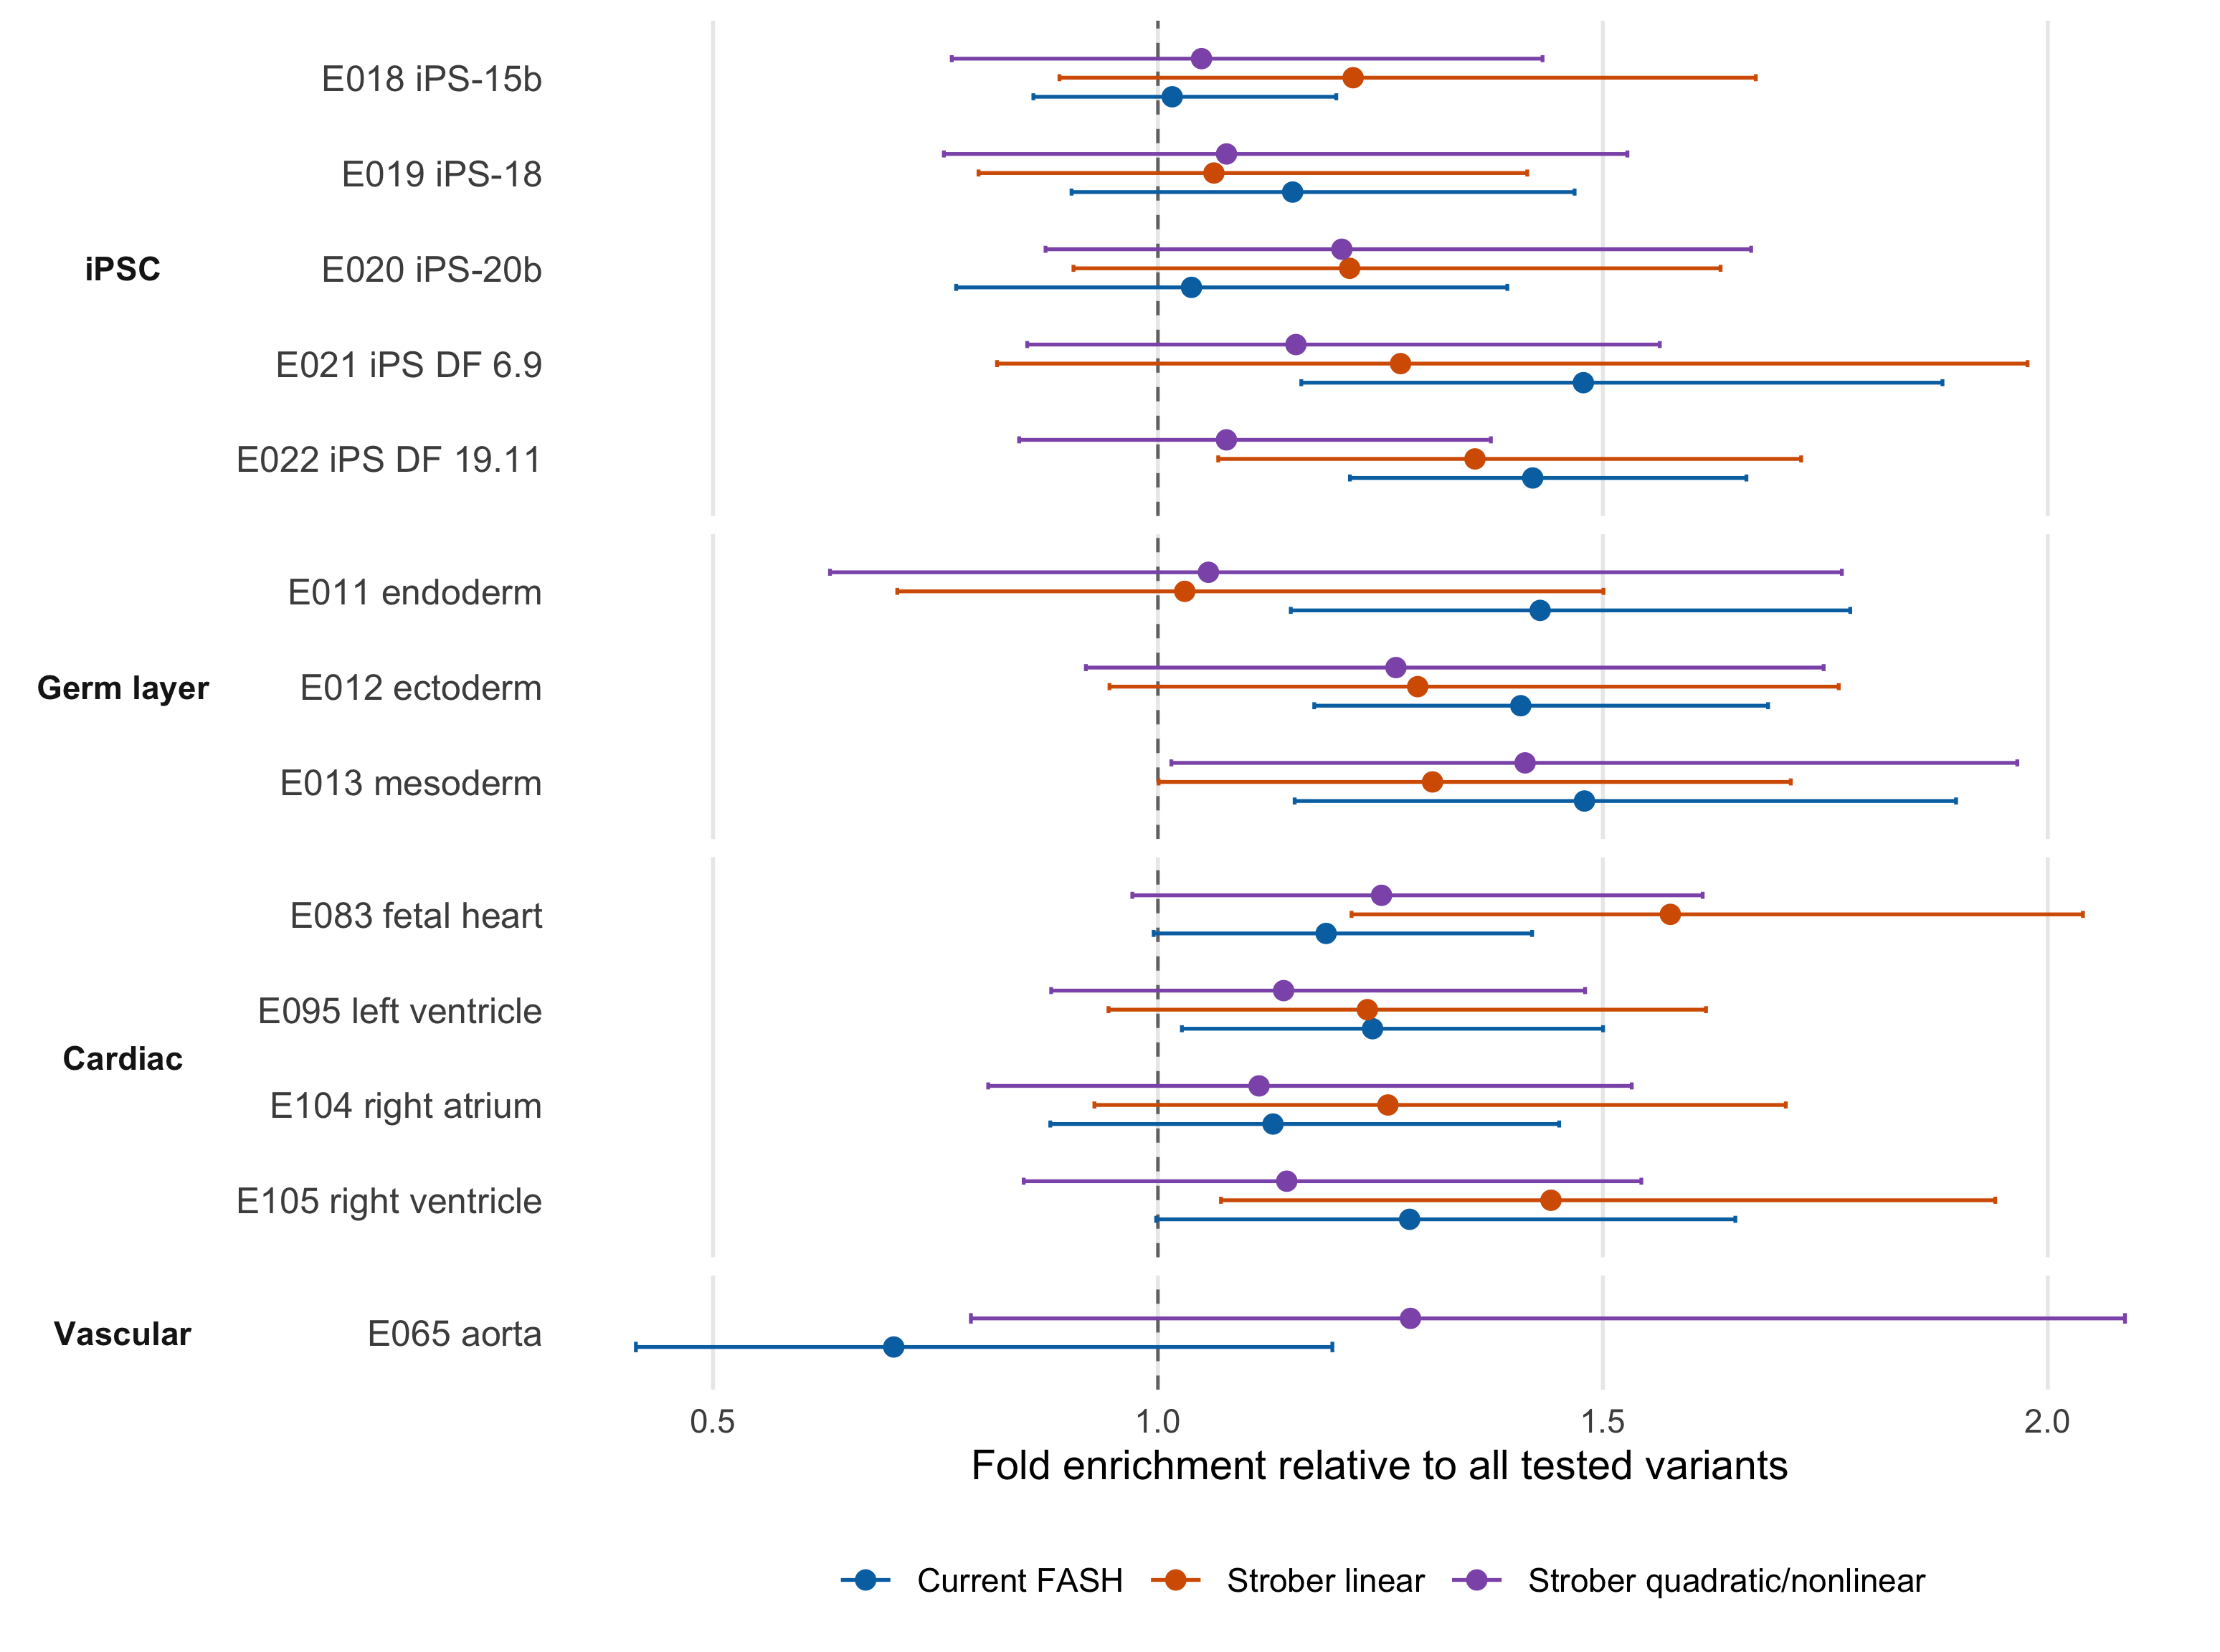
Figure 1. Enhancer enrichment for current FASH, Strober linear, and Strober quadratic/nonlinear discoveries, using one lead variant per discovered gene. Enhancer regions are the Strober et al. (2019) Roadmap 15-state ChromHMM enhancer states, across thirteen epigenomes grouped as iPSC, germ layer, cardiac, and related vascular. Fold enrichment is the overlap rate in the variant set divided by the overlap rate among the remaining tested variants; the dashed line marks no enrichment. Bars are leave-one-autosome-out jackknife 95% intervals. One cell has too few overlapping variants to estimate and is absent: E065 (Strober linear).
FASH’s 1,169 lead variants are enriched in 6 of 13 displayed annotations, over a narrow range of 1.24-1.48-fold. The annotations whose intervals lie entirely above 1 are E021, E022, E011, E012, E013, E095 — that is, two iPSC epigenomes, all three germ layers, and left ventricle. The three remaining cardiac annotations point the same way with intervals that reach or cross 1 (E083 1.19, E105 1.28, E104 1.13).
Both Strober lead sets point the same way. Their intervals are wider, which is what their smaller size would predict — 548 and 690 lead variants against FASH’s 1,169 — and where they do clear 1 it is at annotations drawn from the same groups: E022, E013, E083, E105 for linear and E013 for quadratic/nonlinear. The three discovery sets differ in size, membership and gene composition, so this is not a formal comparison between methods.
The enrichment is spread across pluripotent, germ-layer and cardiac annotations rather than concentrated in any one of them, and within each group some members clear 1 while others do not. These lead-per-gene estimates therefore support the general statement that lead variants identified by FASH are enriched in several established Roadmap enhancer annotations relevant to this differentiation system, and they do not support any claim of developmental-stage or tissue specificity. E065 aorta, the sparsest annotation in the panel and the only related vascular one, carries the single below-1 point estimate, 0.70-fold (95% CI 0.41-1.20), and rests on the fewest overlapping variants, so it should be read as neither support nor contradiction.
Section 2: FASH-specific discoveries
An rsID is counted as FASH-only if it appears in neither Strober analysis, regardless of which gene it was paired with; this is the conservative reading, since an enhancer annotation is a property of a variant rather than of a gene-variant pair. FASH’s lowest-lfdr variant is chosen for each gene first and the exclusion is applied afterwards, so a gene whose best variant Strober also found drops out rather than falling back to a weaker one. That leaves 1,029 of FASH’s 1,169 lead variants.
plot_fash_only()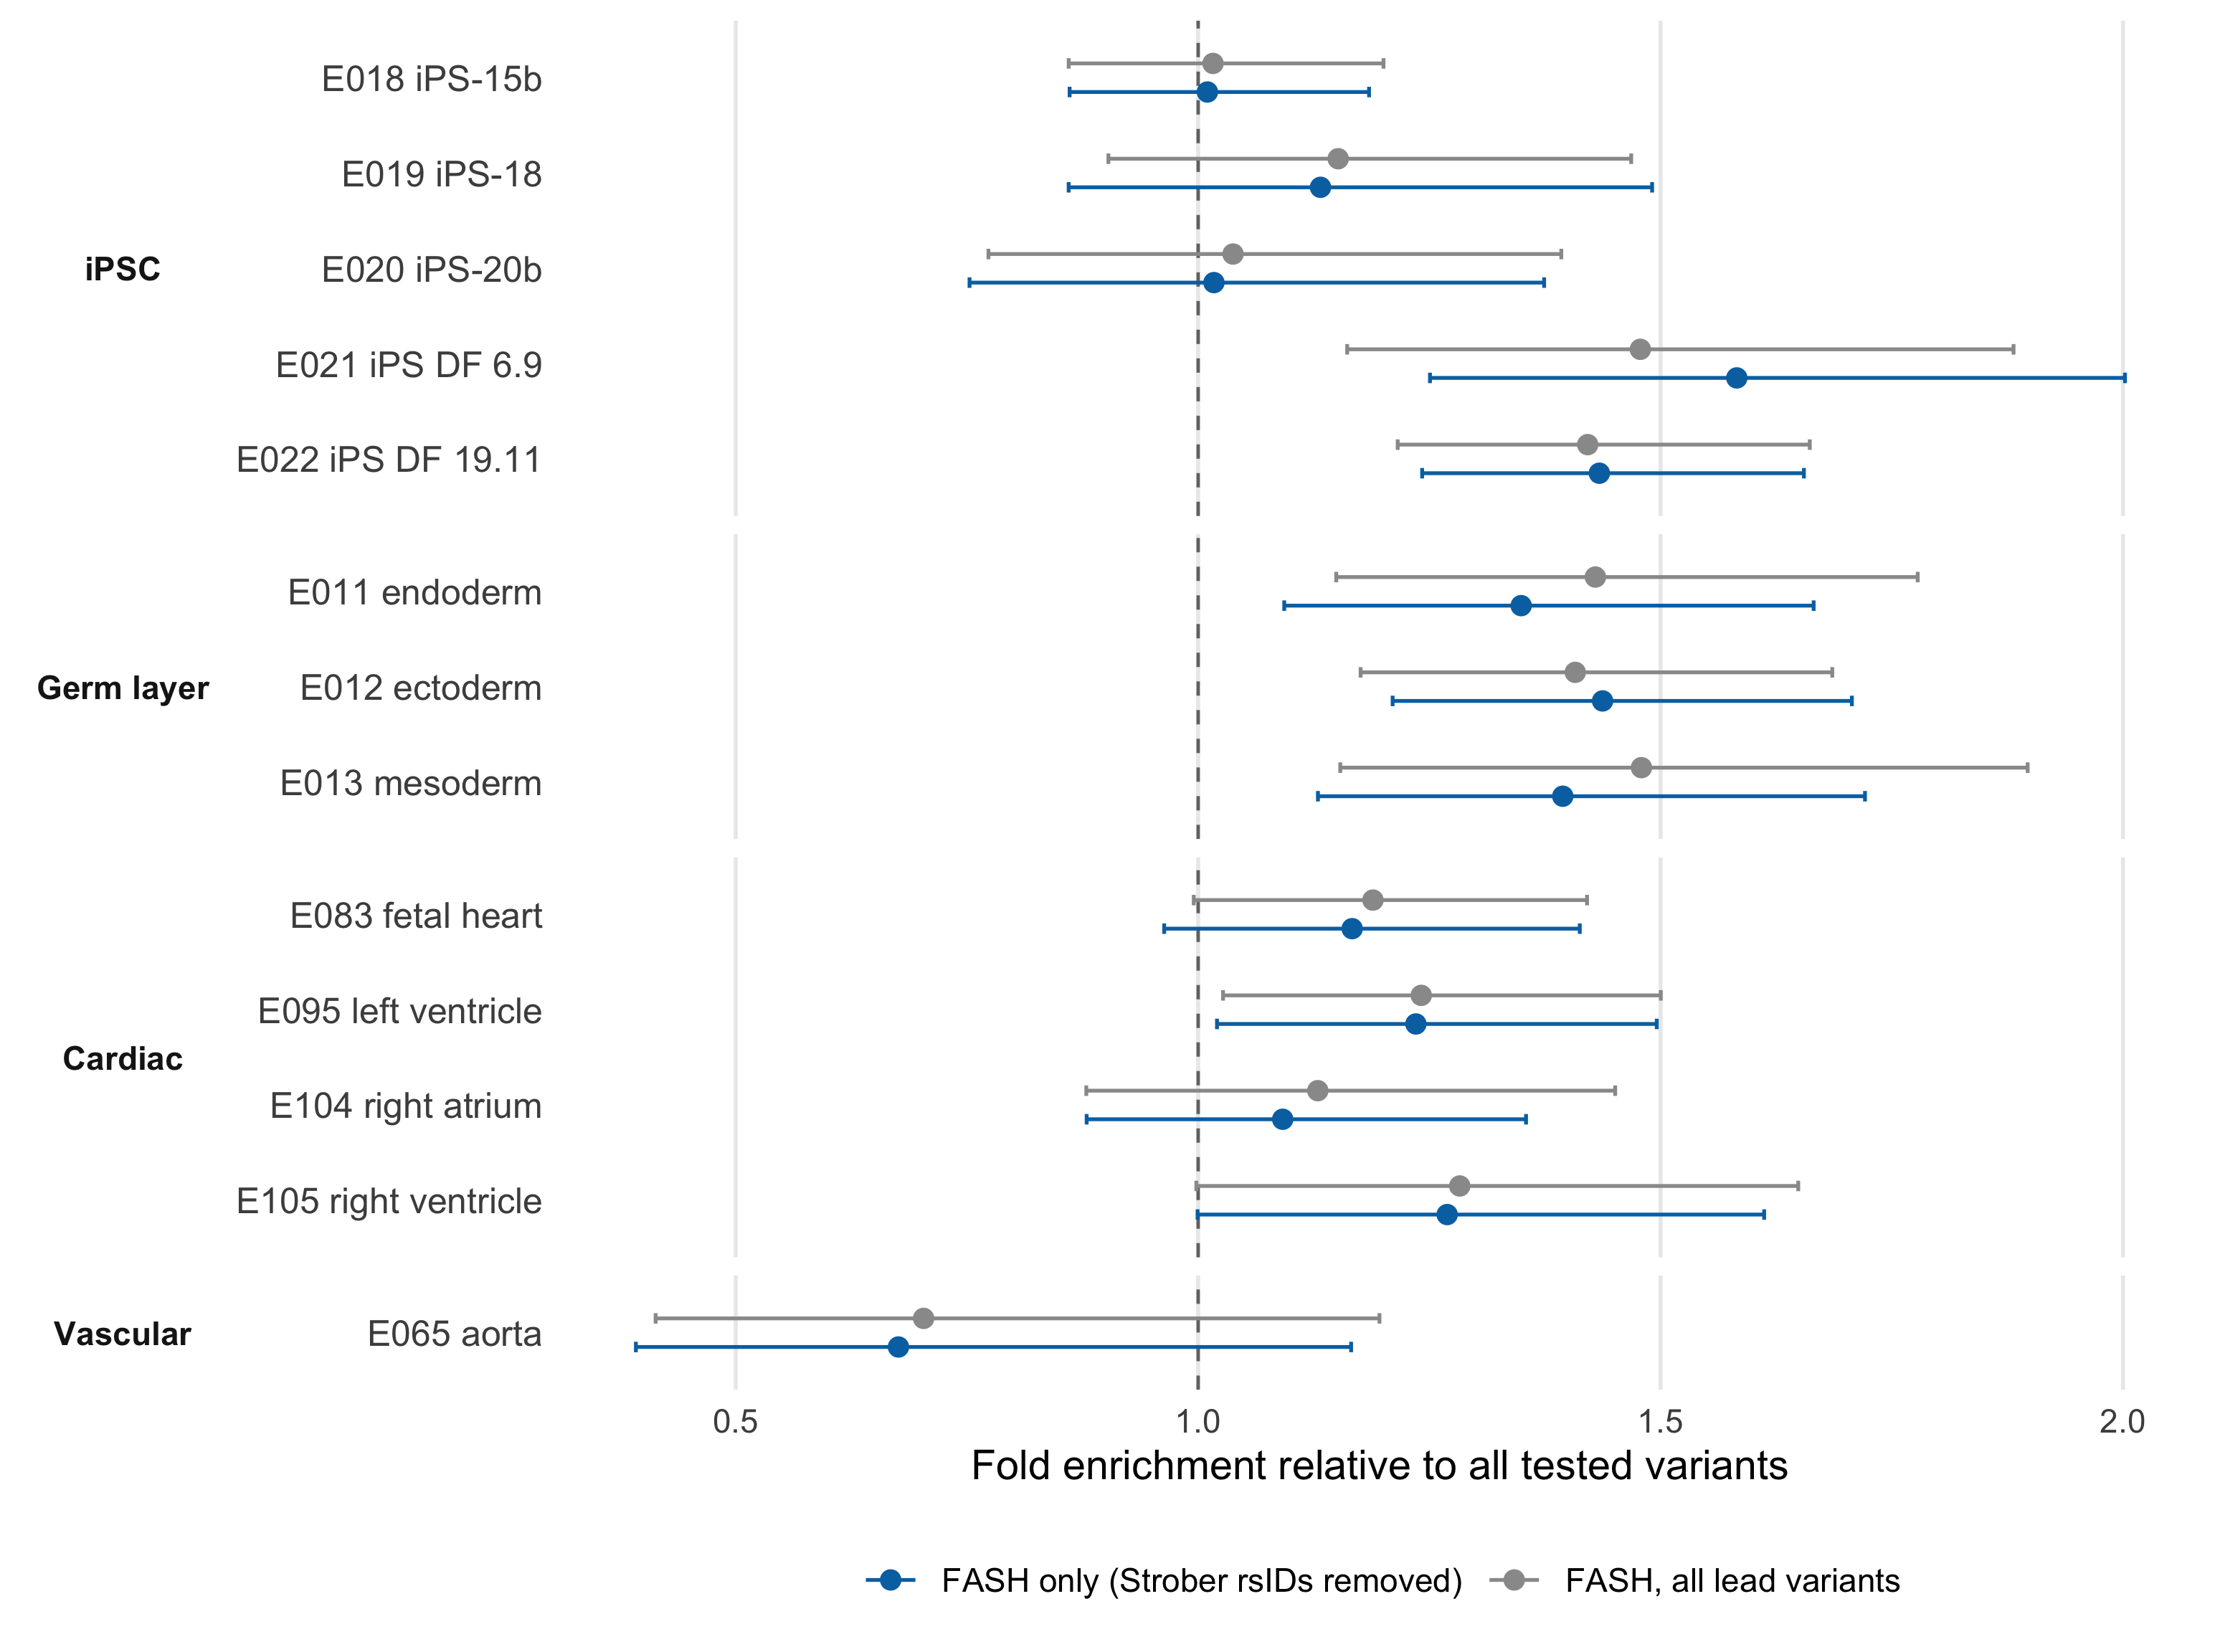
Figure 2. Enhancer enrichment for FASH lead variants at rsIDs that neither Strober analysis identified (blue), against all FASH lead variants as a reference (grey), using one lead variant per discovered gene. Same thirteen Roadmap epigenomes, same Strober et al. (2019) enhancer definition, and same estimator as Figure 1. Bars are leave-one-autosome-out jackknife 95% intervals.
The enrichment survives the exclusion essentially intact. The same 6 of 13 annotations clear 1 — E021, E022, E011, E012, E013, E095 — at 1.24-1.58-fold, against 1.24-1.48-fold for the unfiltered FASH lead set, and the point estimates barely move (E013 1.39 against 1.48; E022 1.43 against 1.42; E095 1.24 against 1.24).
So the enhancer-enrichment pattern is largely retained among FASH lead variants at rsIDs that neither Strober analysis identified. Taken with Section 1, this is orthogonal support for the regulatory relevance of these discoveries as a set, resting on externally established Roadmap and Strober annotations rather than on anything fitted here. It is not validation of any individual discovery, and it does not establish specificity to a particular differentiation stage or tissue.
Methods
Background. All 745,867 variants tested in this study that lie on an autosome and participate in at least one tested gene-variant pair. No external reference panel enters, so the background is exactly the cis-eQTL testing universe from which the discoveries were drawn.
Discovery sets and lead variants. FASH uses
cumulative lfdr at FDR 0.05 after the BF correction; both Strober
analyses use their reported eFDR 0.05, with
non_linear_dynamic_eqtls_5_pc.txt labelled
quadratic/nonlinear throughout. Every set displayed here keeps, for each
discovered gene, one lead variant: the variant with the lowest lfdr
(FASH) or the smallest p-value (Strober). Using one lead variant per
gene reduces the repeated contribution of multiple correlated variants
associated with the same gene.
Enhancer annotation. Roadmap Epigenomics 15-state
ChromHMM segmentations for the thirteen epigenomes above, with enhancer
regions defined by the states Enh, EnhG,
EnhBiv and BivFlnk as in Strober et
al. (2019). The iPSC and heart epigenome choices follow the same
source.
Fold enrichment. For a variant set and an enhancer annotation,
\[\text{fold enrichment} = \frac{\text{proportion of selected variants overlapping the enhancer}}{\text{proportion of the remaining tested variants overlapping the enhancer}}\]
The background excludes the set being tested, so a set is never part of its own comparator. No matched controls are used: the purpose here is a simple exploratory check, and every background variant already comes from the same testing universe. A set-annotation pair with fewer than 10 overlapping variants is reported as not estimable rather than estimated.
Uncertainty. Uncertainty was assessed using a leave-one-autosome-out jackknife to account approximately for local dependence among variants. All intervals shown are 95% jackknife intervals; nominal p-values are available in the output CSVs but are not used on this page.
Reproducibility
The figures on this page read the validated thirteen-epigenome
indicator table and estimates built by run_roadmap_enhancer_exploration.R,
through the shared estimator in enrichment_api.R.
The annotation intervals are not rebuilt here. Exactly the calls below
reproduce every estimate shown above:
source(
  "code/revision_simulations/internal/fash_strober_enhancer_comparison/enrichment_api.R"
)
source(
  "code/revision_simulations/internal/roadmap_enhancer_exploration/roadmap_enhancer_exploration_helpers.R"
)

# 1. Background: every autosomal variant tested in this study.
background <- load_background()

# 2. Discovery sets, one lead variant per discovered gene.
sets <- load_variant_sets()[c(
  "current_lead", "linear_lead", "quadratic_lead", "current_only_lead"
)]

# 3. The thirteen Roadmap epigenomes, as the indicator table built by the
#    exploration run script.
annotations <- readRDS(file.path(
  roadmap_output_directory(), "roadmap_enhancer_annotation_matrix.rds"
))

# 4. The Strober four-state enhancer definition, one column per epigenome.
strober <- grep(
  "[Strober EnhG+Enh+EnhBiv+BivFlnk]", names(annotations),
  fixed = TRUE, value = TRUE
)

# 5. Fold enrichment against the tested-variant background, with
# 6. leave-one-autosome-out jackknife 95% intervals.
compute_enrichment(
  sets,
  background,
  annotations[, c("variant_id", "chromosome", strober)],
  controls = "background"
)
compute_enrichment() takes any named list of variant
vectors, so substituting your own sets needs no other change:
compute_enrichment(
  list(my_hits = readLines("my_variants.txt"), fash_only = sets$current_only_lead),
  background,
  annotations[, c("variant_id", "chromosome", strober)],
  controls = "background"
)
Rebuilding the inputs from source — the tested variant universe, then the discovery sets, then the thirteen Roadmap segmentations and the estimates this page displays:
Rscript --vanilla \
  code/revision_simulations/internal/variant_annotation_enrichment/download_variant_annotations.R
Rscript --vanilla \
  code/revision_simulations/internal/variant_annotation_enrichment/run_variant_annotation_enrichment.R
Rscript --vanilla \
  code/revision_simulations/internal/fash_strober_enhancer_comparison/run_fash_strober_enhancer_comparison.R
Rscript --vanilla \
  code/revision_simulations/internal/roadmap_enhancer_exploration/run_roadmap_enhancer_exploration.R
Intermediate data stays under output/, which is not
tracked in git; the repository carries code, not data. Rebuilding this
page:
Rscript --vanilla -e \
  'workflowr::wflow_build("analysis/revision_enhancer_enrichment.rmd")'
References
Roadmap Epigenomics Consortium, Kundaje, A., Meuleman, W., Ernst, J., Bilenky, M., Yen, A., et al. (2015). Integrative analysis of 111 reference human epigenomes. Nature 518, 317–330. https://doi.org/10.1038/nature14248
Strober, B., Elorbany, R., Rhodes, K., Krishnan, N., Tayeb, K., Battle, A., and Gilad, Y. (2019). Dynamic genetic regulation of gene expression during cellular differentiation. Science 364, 1287–1290. https://doi.org/10.1126/science.aaw0040
sessionInfo()R version 4.5.1 (2025-06-13)
Platform: aarch64-apple-darwin20
Running under: macOS Sequoia 15.6.1
Matrix products: default
BLAS: /System/Library/Frameworks/Accelerate.framework/Versions/A/Frameworks/vecLib.framework/Versions/A/libBLAS.dylib
LAPACK: /Library/Frameworks/R.framework/Versions/4.5-arm64/Resources/lib/libRlapack.dylib; LAPACK version 3.12.1
locale:
[1] en_CA/en_CA/en_CA/C/en_CA/en_CA
time zone: America/Chicago
tzcode source: internal
attached base packages:
[1] stats graphics grDevices utils datasets methods base
loaded via a namespace (and not attached):
[1] gtable_0.3.6 jsonlite_2.0.0 dplyr_1.1.4 compiler_4.5.1
[5] promises_1.5.0 tidyselect_1.2.1 Rcpp_1.1.1-1.1 stringr_1.6.0
[9] git2r_0.36.2 dichromat_2.0-0.1 later_1.4.4 jquerylib_0.1.4
[13] scales_1.4.0 yaml_2.3.12 fastmap_1.2.0 ggplot2_4.0.3
[17] R6_2.6.1 labeling_0.4.3 generics_0.1.4 workflowr_1.7.2
[21] knitr_1.50 tibble_3.3.0 rprojroot_2.1.1 bslib_0.9.0
[25] pillar_1.11.1 RColorBrewer_1.1-3 rlang_1.2.0 cachem_1.1.0
[29] stringi_1.8.9 httpuv_1.6.16 xfun_0.55 S7_0.2.2
[33] fs_1.6.6 sass_0.4.10 otel_0.2.0 cli_3.6.6
[37] withr_3.0.3 magrittr_2.0.5 digest_0.6.39 grid_4.5.1
[41] rstudioapi_0.17.1 lifecycle_1.0.5 vctrs_0.7.3 evaluate_1.0.5
[45] glue_1.8.1 farver_2.1.2 whisker_0.4.1 rmarkdown_2.30
[49] tools_4.5.1 pkgconfig_2.0.3 htmltools_0.5.9