Figure S2: GP30 and GP58 loadings across T-cell subsets
Ziang Zhang
Last updated: 2026-07-02
Checks: 5 1
Knit directory:
immgenT-GP-analysis/analysis/
This reproducible R Markdown analysis was created with workflowr (version 1.7.2). The Checks tab describes the reproducibility checks that were applied when the results were created. The Past versions tab lists the development history.
Great job! The global environment was empty. Objects defined in the global environment can affect the analysis in your R Markdown file in unknown ways. For reproduciblity it’s best to always run the code in an empty environment.
The command set.seed(1) was run prior to running the
code in the R Markdown file. Setting a seed ensures that any results
that rely on randomness, e.g. subsampling or permutations, are
reproducible.
Great job! Recording the operating system, R version, and package versions is critical for reproducibility.
Nice! There were no cached chunks for this analysis, so you can be confident that you successfully produced the results during this run.
Great job! Using relative paths to the files within your workflowr project makes it easier to run your code on other machines.
Tracking code development and connecting the code version to the
results is critical for reproducibility. To start using Git, open the
Terminal and type git init in your project directory.
This project is not being versioned with Git. To obtain the full
reproducibility benefits of using workflowr, please see
?wflow_start.
All panels are produced by script-refactor/FigureS2.R,
which shares its source data setup with Figure
2. The code below is shown for reference (not re-executed on this
page); the images are its pre-rendered output.
Setup
# Figure S2. GP30 and GP58 loadings across T-cell subsets.
#
# Panels produced (see figures/final-selected/bits/Figure S2/FigureS2_caption.md
# for the full caption text):
#   S2A  Boxplots of GP30 loading across Tz subsets (iNKT, MAIT, other Tz)
#        vs all other, non-Tz T cells.
#   S2B  Boxplots of GP58 loading: resting CD8, activated CD8, vs the other
#        six T-cell lineages pooled as "Other T cells".
#   S2C  Boxplots of CD8A/CD8B log-normalized CITE-seq protein expression in
#        CD8 cells, resting vs activated.
#
# Source: ported from script/Figure_Lineage.R (see Figure2.R for the main
# Figure 2 panels from the same file).

library(ggplot2)
library(dplyr)
library(ggrastr)
library(tidyr)
library(Matrix) # protein_mat_normalized_lognorm is a dgCMatrix; rownames()
                # dispatch on it is unreliable unless Matrix is attached
                # (not just loaded as a namespace), which silently intersected
                # to zero cells during this refactor -- keep this library() call.

data_path <- "data/"
figure_path <- "figure-refactor/Figure S2/"
source("code-refactor/R/plot_utils.R") # tukey_outliers()

# ============================================================
# Load data
# ============================================================
seurat_meta <- readRDS(paste0(data_path, "igt1_96_withtotalvi20260206_clean_ADTonly.Rds"))@meta.data
L_pm_filtered <- readRDS(paste0(data_path, "L_pm_filtered.rds"))
colnames(L_pm_filtered) <- gsub("^K", "GP", colnames(L_pm_filtered))
seurat_meta_filtered <- seurat_meta[rownames(L_pm_filtered), ]
selected_lineage_in_order <- c("CD8", "CD4", "Treg", "gdT", "CD8aa", "Tz", "DN")
(A) GP30 loading across Tz subsets
# ============================================================
# S2A: GP30 loading, Tz subsets (iNKT/MAIT/other Tz) vs other T cells
# ============================================================
GP30_df <- seurat_meta_filtered %>%
  select(cellID, annotation_level1, iNKT, MAIT) %>%
  filter(annotation_level1 != "thymocyte") %>%
  mutate(GP30_loading = L_pm_filtered[cellID, "GP30"]) %>%
  mutate(
    Group = case_when(
      annotation_level1 == "Tz" & iNKT ~ "iNKT",
      annotation_level1 == "Tz" & MAIT ~ "MAIT",
      annotation_level1 == "Tz" ~ "Other Tz",
      TRUE ~ "Other T cells"
    ),
    Group = factor(Group, levels = c("iNKT", "MAIT", "Other Tz", "Other T cells"))
  )

p_S2A <- ggplot(GP30_df, aes(x = Group, y = GP30_loading, fill = Group)) +
  geom_boxplot(outlier.shape = NA) +
  ggrastr::rasterise(
    geom_point(data = tukey_outliers(GP30_df, "GP30_loading", "Group"), size = 0.5, alpha = 0.3, show.legend = FALSE),
    dpi = 300
  ) +
  scale_fill_manual(values = c("iNKT" = "darkgoldenrod2", "MAIT" = "darkgoldenrod3", "Other Tz" = "darkgoldenrod1", "Other T cells" = "grey70")) +
  labs(title = "GP30 loading across Tz subsets and other T cells", x = NULL, y = "GP30 loading") +
  theme_minimal(base_size = 12) +
  theme(plot.title = element_text(face = "bold", hjust = 0.5), panel.grid.major.x = element_blank(), legend.position = "none")
ggsave(filename = paste0(figure_path, "S2A.pdf"), plot = p_S2A, width = 6, height = 5)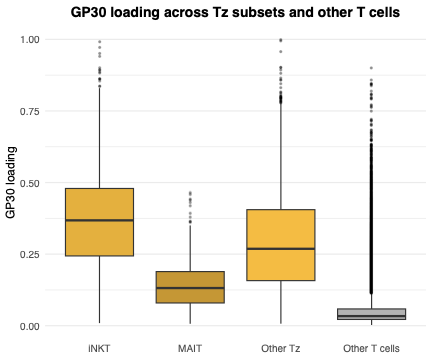
Fig. S2A. Boxplots of GP30 loading across Tz subsets – iNKT, MAIT, and other Tz (Tz cells that are neither iNKT nor MAIT) – and all other, non-Tz T cells; thymocytes are excluded. Boxes show the median and interquartile range, whiskers extend to 1.5x the interquartile range, and points mark values beyond the whiskers.
(B) GP58 loading, resting/activated CD8
# ============================================================
# S2B: GP58 loading, resting/activated CD8 vs other T cells
# ============================================================
non_CD8_lineages <- setdiff(selected_lineage_in_order, "CD8")
GP58_df <- seurat_meta_filtered %>%
  select(cellID, annotation_level1, annotation_level2_group) %>%
  mutate(GP58_loading = L_pm_filtered[cellID, "GP58"]) %>%
  mutate(
    Group = case_when(
      annotation_level1 == "CD8" & annotation_level2_group == "resting" ~ "CD8 (Resting)",
      annotation_level1 == "CD8" & annotation_level2_group == "activated" ~ "CD8 (Activated)",
      annotation_level1 %in% non_CD8_lineages ~ "Other T cells",
      TRUE ~ NA_character_
    )
  ) %>%
  filter(!is.na(Group)) %>%
  mutate(Group = factor(Group, levels = c("CD8 (Resting)", "CD8 (Activated)", "Other T cells")))

p_S2B <- ggplot(GP58_df, aes(x = Group, y = GP58_loading, fill = Group)) +
  geom_boxplot(outlier.shape = NA) +
  ggrastr::rasterise(
    geom_point(data = tukey_outliers(GP58_df, "GP58_loading", "Group"), size = 0.5, alpha = 0.3, show.legend = FALSE),
    dpi = 300
  ) +
  scale_fill_manual(values = c("CD8 (Resting)" = "darkorange2", "CD8 (Activated)" = "orange", "Other T cells" = "grey70")) +
  labs(title = "GP58 loading across CD8 subsets and other T cells", x = NULL, y = "GP58 loading") +
  theme_minimal(base_size = 12) +
  theme(plot.title = element_text(face = "bold", hjust = 0.5), panel.grid.major.x = element_blank(), legend.position = "none")
ggsave(filename = paste0(figure_path, "S2B.pdf"), plot = p_S2B, width = 5, height = 5)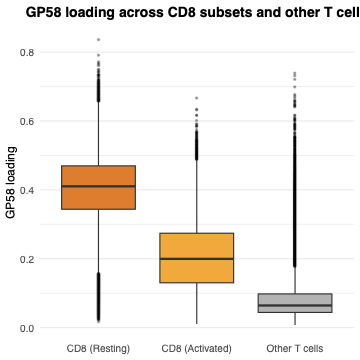
Fig. S2B. Boxplots of GP58 loading in resting CD8, activated CD8 (split by activation annotation), and the other six T-cell lineages pooled as “Other T cells” (CD4, Treg, gdT, CD8aa, Tz, and DN; thymocytes and DP cells are excluded); boxes and outlier points as in (A).
(C) CD8A/CD8B protein expression
# ============================================================
# S2C: CD8A/CD8B CITE-seq protein expression, resting vs activated CD8
# ============================================================
protein_mat_normalized_lognorm <- readRDS(paste0(data_path, "protein_mat_normalized_lognorm.rds"))

CD8_citeseq_cells <- seurat_meta_filtered$cellID[
  seurat_meta_filtered$annotation_level1 == "CD8" &
    seurat_meta_filtered$cite_seq &
    seurat_meta_filtered$annotation_level2_group %in% c("resting", "activated")
]
CD8_citeseq_cells <- intersect(CD8_citeseq_cells, rownames(protein_mat_normalized_lognorm))

CD8_protein_df <- data.frame(
  cellID = CD8_citeseq_cells,
  Group = ifelse(seurat_meta_filtered[CD8_citeseq_cells, "annotation_level2_group"] == "resting", "CD8 (Resting)", "CD8 (Activated)"),
  CD8A = protein_mat_normalized_lognorm[CD8_citeseq_cells, "CD8A"],
  CD8B = protein_mat_normalized_lognorm[CD8_citeseq_cells, "CD8B"]
) %>%
  tidyr::pivot_longer(cols = c("CD8A", "CD8B"), names_to = "Protein", values_to = "Expression") %>%
  mutate(Group = factor(Group, levels = c("CD8 (Resting)", "CD8 (Activated)")))

p_S2C <- ggplot(CD8_protein_df, aes(x = Protein, y = Expression, fill = Group)) +
  geom_boxplot(outlier.shape = NA) +
  ggrastr::rasterise(
    geom_point(
      data = tukey_outliers(CD8_protein_df, "Expression", c("Protein", "Group")),
      aes(group = Group), size = 0.5, alpha = 0.3, position = position_dodge(width = 0.75), show.legend = FALSE
    ),
    dpi = 300
  ) +
  scale_fill_manual(values = c("CD8 (Resting)" = "darkorange2", "CD8 (Activated)" = "orange")) +
  labs(title = "CD8A / CD8B protein expression in CD8 cells", x = NULL, y = "Log-normalized protein expression", fill = NULL) +
  theme_minimal(base_size = 12) +
  theme(plot.title = element_text(face = "bold", hjust = 0.5), panel.grid.major.x = element_blank(), legend.position = "top")
ggsave(filename = paste0(figure_path, "S2C.pdf"), plot = p_S2C, width = 5, height = 5)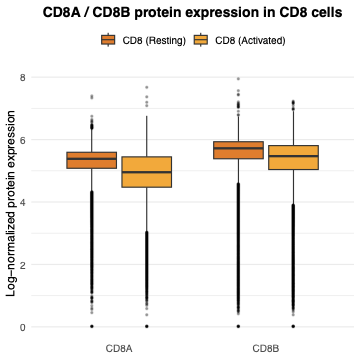
Fig. S2C. Boxplots of CD8A and CD8B log-normalized surface-protein (CITE-seq) expression in CD8 cells, comparing resting versus activated, restricted to CD8 cells with CITE-seq protein measurements; boxes and outlier points as in (A).
sessionInfo()R version 4.5.1 (2025-06-13)
Platform: aarch64-apple-darwin20
Running under: macOS Sequoia 15.6.1
Matrix products: default
BLAS: /System/Library/Frameworks/Accelerate.framework/Versions/A/Frameworks/vecLib.framework/Versions/A/libBLAS.dylib
LAPACK: /Library/Frameworks/R.framework/Versions/4.5-arm64/Resources/lib/libRlapack.dylib; LAPACK version 3.12.1
locale:
[1] en_CA/en_CA/en_CA/C/en_CA/en_CA
time zone: Asia/Tokyo
tzcode source: internal
attached base packages:
[1] stats graphics grDevices utils datasets methods base
loaded via a namespace (and not attached):
[1] vctrs_0.7.3 cli_3.6.6 knitr_1.50 rlang_1.2.0
[5] xfun_0.55 stringi_1.8.7 otel_0.2.0 promises_1.5.0
[9] jsonlite_2.0.0 workflowr_1.7.2 glue_1.8.1 rprojroot_2.1.1
[13] git2r_0.36.2 htmltools_0.5.9 httpuv_1.6.16 sass_0.4.10
[17] rmarkdown_2.30 evaluate_1.0.5 jquerylib_0.1.4 tibble_3.3.0
[21] fastmap_1.2.0 yaml_2.3.12 lifecycle_1.0.5 stringr_1.6.0
[25] compiler_4.5.1 fs_1.6.6 Rcpp_1.1.1-1.1 pkgconfig_2.0.3
[29] later_1.4.4 digest_0.6.39 R6_2.6.1 pillar_1.11.1
[33] magrittr_2.0.5 bslib_0.9.0 tools_4.5.1 cachem_1.1.0