Figure 3: GPs associated with T-cell activation
Ziang Zhang
Last updated: 2026-07-03
Checks: 7 0
Knit directory:
immgenT-GP-analysis/analysis/
This reproducible R Markdown analysis was created with workflowr (version 1.7.2). The Checks tab describes the reproducibility checks that were applied when the results were created. The Past versions tab lists the development history.
Great! Since the R Markdown file has been committed to the Git repository, you know the exact version of the code that produced these results.
Great job! The global environment was empty. Objects defined in the global environment can affect the analysis in your R Markdown file in unknown ways. For reproduciblity it’s best to always run the code in an empty environment.
The command set.seed(1) was run prior to running the
code in the R Markdown file. Setting a seed ensures that any results
that rely on randomness, e.g. subsampling or permutations, are
reproducible.
Great job! Recording the operating system, R version, and package versions is critical for reproducibility.
Nice! There were no cached chunks for this analysis, so you can be confident that you successfully produced the results during this run.
Great job! Using relative paths to the files within your workflowr project makes it easier to run your code on other machines.
Great! You are using Git for version control. Tracking code development and connecting the code version to the results is critical for reproducibility.
The results in this page were generated with repository version db7a5cd. See the Past versions tab to see a history of the changes made to the R Markdown and HTML files.
Note that you need to be careful to ensure that all relevant files for
the analysis have been committed to Git prior to generating the results
(you can use wflow_publish or
wflow_git_commit). workflowr only checks the R Markdown
file, but you know if there are other scripts or data files that it
depends on. Below is the status of the Git repository when the results
were generated:
Ignored files:
    Ignored:    analysis/.DS_Store
    Ignored:    analysis/.Rhistory
    Ignored:    analysis/assets/.DS_Store
    Ignored:    code 2/
    Ignored:    code/pipeline 2/
    Ignored:    data
    Ignored:    figures/final-selected/.DS_Store
    Ignored:    figures/final-selected/bits/.DS_Store
    Ignored:    figures/final-selected/bits/Figure 1/.DS_Store
    Ignored:    figures/final-selected/bits/Figure 2/.DS_Store
    Ignored:    figures/final-selected/bits/Figure 3/.DS_Store
    Ignored:    figures/final-selected/bits/Figure 4/.DS_Store
    Ignored:    figures/final-selected/bits/Figure 6/.DS_Store
    Ignored:    figures/final-selected/bits/Figure 7/.DS_Store
    Ignored:    figures/final-selected/bits/Figure S1/.DS_Store
    Ignored:    figures/final-selected/bits/Figure S2/.DS_Store
    Ignored:    figures/final-selected/bits/Figure S3/.DS_Store
    Ignored:    figures/final-selected/bits/Figure S6/.DS_Store
    Ignored:    figures/final-selected/bits/Figure S7/.DS_Store
    Ignored:    figures/generated/Figure 1 2/
    Ignored:    figures/generated/Figure 5/
    Ignored:    figures/generated/Figure 7/
    Ignored:    figures/generated/Figure 8/
    Ignored:    figures/generated/Figure 9/
    Ignored:    figures/generated/Figure S1 2/
    Ignored:    figures/generated/Figure S2 2/
    Ignored:    figures/generated/Figure S3 2/
    Ignored:    figures/generated/Figure S6 2/

Untracked files:
    Untracked:  code/pipeline/01b_filter_cells.R

Unstaged changes:
    Modified:   code/README.md
    Modified:   code/pipeline/02_compute_auc.R
    Modified:   code/pipeline/04_protein_projection.R
    Modified:   figures/generated/Figure 1/1A.pdf
    Modified:   figures/generated/Figure 1/1C.pdf
    Modified:   figures/generated/Figure 1/1D.pdf
    Modified:   figures/generated/Figure 1/1E.pdf
    Modified:   figures/generated/Figure 1/1F.pdf
    Modified:   figures/generated/Figure 1/1G.pdf
    Modified:   figures/generated/Figure 1/1H.pdf
    Modified:   figures/generated/Figure 1/1I.pdf
    Modified:   figures/generated/Figure 1/hist_active_cells_per_GP.pdf
    Modified:   figures/generated/Figure 1/hist_active_genes_prop_per_GP.pdf
    Modified:   figures/generated/Figure 1/scatter_e_active_cells_vs_genes.pdf
    Modified:   figures/generated/Figure 2/2A.pdf
    Modified:   figures/generated/Figure 2/2B.pdf
    Modified:   figures/generated/Figure 2/2C.pdf
    Modified:   figures/generated/Figure 2/2D.pdf
    Modified:   figures/generated/Figure 2/2E.pdf
    Modified:   figures/generated/Figure 2/2F.pdf
    Modified:   figures/generated/Figure 2/2G.pdf
    Modified:   figures/generated/Figure 2/2H.pdf
    Modified:   figures/generated/Figure 2/2I.pdf
    Modified:   figures/generated/Figure 2/2J.pdf
    Modified:   figures/generated/Figure 2/2K.pdf
    Modified:   figures/generated/Figure 2/2L.pdf
    Modified:   figures/generated/Figure 2/2M.pdf
    Modified:   figures/generated/Figure 3/3c.pdf
    Modified:   figures/generated/Figure 3/3d.pdf
    Modified:   figures/generated/Figure 3/3e.pdf
    Modified:   figures/generated/Figure 3/3f.pdf
    Modified:   figures/generated/Figure 3/3g.pdf
    Modified:   figures/generated/Figure 4/4a.pdf
    Modified:   figures/generated/Figure 4/4b.pdf
    Modified:   figures/generated/Figure 4/4c.pdf
    Modified:   figures/generated/Figure 4/4d.pdf
    Modified:   figures/generated/Figure 4/4e.pdf
    Modified:   figures/generated/Figure 6/6b.pdf
    Modified:   figures/generated/Figure 6/6c.pdf
    Modified:   figures/generated/Figure 6/6d.pdf
    Modified:   figures/generated/Figure 6/6e.pdf
    Modified:   figures/generated/Figure 6/6f.pdf
    Modified:   figures/generated/Figure 6/6g.pdf
    Modified:   figures/generated/Figure 6/6h.pdf
    Modified:   figures/generated/Figure 6/6i.pdf
    Modified:   figures/generated/Figure 6/6j.pdf
    Modified:   figures/generated/Figure 6/6k.pdf
    Modified:   figures/generated/Figure S1/S1A.pdf
    Modified:   figures/generated/Figure S1/S1B.pdf
    Modified:   figures/generated/Figure S1/S1C.pdf
    Modified:   figures/generated/Figure S1/S1D.pdf
    Modified:   figures/generated/Figure S2/S2A.pdf
    Modified:   figures/generated/Figure S2/S2B.pdf
    Modified:   figures/generated/Figure S2/S2C.pdf
    Modified:   figures/generated/Figure S3/s3c.pdf
    Modified:   figures/generated/Figure S3/s3d.pdf
    Modified:   figures/generated/Figure S3/s3e.pdf
    Modified:   figures/generated/Figure S3/s3f.pdf
    Modified:   figures/generated/Figure S3/s3g.pdf
    Modified:   figures/generated/Figure S3/s3h.pdf
    Modified:   script/Figure1.R
    Modified:   script/Figure2.R
    Modified:   script/Figure3.R
    Modified:   script/Figure4.R
    Modified:   script/Figure6.R
    Modified:   script/FigureS1.R
    Modified:   script/FigureS2.R
    Modified:   script/FigureS3.R
    Modified:   script/FigureS6.R
    Modified:   script/TableS1.R
Note that any generated files, e.g. HTML, png, CSS, etc., are not included in this status report because it is ok for generated content to have uncommitted changes.
These are the previous versions of the repository in which changes were
made to the R Markdown (analysis/Figure3.Rmd) and HTML
(docs/Figure3.html) files. If you’ve configured a remote
Git repository (see ?wflow_git_remote), click on the
hyperlinks in the table below to view the files as they were in that
past version.
| File | Version | Author | Date | Message |
|---|---|---|---|---|
| html | 827c89b | Ziang Zhang | 2026-07-03 | Build site. |
| Rmd | 8ac7f9f | Ziang Zhang | 2026-07-03 | Add data provenance notes to each script; remove conversational |
| Rmd | f9db962 | Ziang Zhang | 2026-07-02 | Simplify layout: drop old code/script folders, rename |
| html | c6e5086 | Ziang Zhang | 2026-07-02 | Build site. |
| html | 5a79883 | Ziang Zhang | 2026-07-02 | Build site. |
| Rmd | 2b0e445 | Ziang Zhang | 2026-07-02 | Fix GitHub source links to point at the new |
| html | cf1d0ac | Ziang Zhang | 2026-07-02 | Build site. |
| Rmd | 06b2461 | Ziang Zhang | 2026-07-02 | Initial commit: immgenT-GP-analysis |
| html | 06b2461 | Ziang Zhang | 2026-07-02 | Initial commit: immgenT-GP-analysis |
All panels are produced by script/Figure3.R,
which shares its curated GP set with Figure
S3 via code/R/activation_shared_setup.R. The code below
is shown for reference (not re-executed on this page); the images are
its pre-rendered output.
Setup
Data loading, shared across all panels below.
# Figure 3. GPs associated with T-cell activation.
#
# Panels produced (see figures/Figure3_Activation/Figure3_caption.md for the
# full caption text -- lettered a-e there, but relettered c-g in the final
# figures/final-selected/bits/Figure 3/ bundle; this script uses the final
# c-g lettering to match):
#   3c  Standardized mean difference (d) in GP loading, activated vs resting,
#       CD4 (x) vs CD8 (y); curated GPs colored by semantic group and labeled.
#   3d  GP-gene signature network: each curated GP linked to its top 5
#       positively/negatively regulated genes.
#   3e  Bipartite TF-GP network for the curated activation GPs.
#   3f  Heatmap of log2FC in mean GP loading across experimental conditions,
#       for activated CD4/CD8 cells.
#   3g  Heatmap of mean GP loading per Level-2 sub-lineage, across the 7
#       T-cell lineages.
#
# Source: ported from Figure_Activation.R, which also produced the
# Figure S3 panels (see FigureS3.R) from the same curated GP set and cell
# groupings -- that shared setup now lives in
# code/R/activation_shared_setup.R, sourced by both scripts.
#
# Required inputs (data/) -- see code/README.md's "Data provenance" table
# for the full picture:
#   L_pm_filtered.rds, F_pm_filtered.rds     [code/pipeline/01b_filter_cells.R]
#   igt1_96_..._ADTonly.Rds                  [primary input Seurat object]

library(ggplot2)
library(ggrepel)
library(dplyr)
library(tidygraph)
library(ggraph)
library(pheatmap)
library(scales)

data_path <- "data/"
figure_path <- "figures/generated/Figure 3/"
source("code/R/plot_utils.R") # scale_cols()
source("code/R/tf_network.R") # optimize_bipartite_order(), plot_tf_gp_network_v2()

# ============================================================
# Load data
# ============================================================
L_pm_filtered <- readRDS(paste0(data_path, "L_pm_filtered.rds"))
colnames(L_pm_filtered) <- paste0("GP", seq_len(ncol(L_pm_filtered)))
F_pm_filtered <- readRDS(paste0(data_path, "F_pm_filtered.rds"))
colnames(F_pm_filtered) <- paste0("GP", seq_len(ncol(F_pm_filtered)))
seurat_meta <- readRDS(paste0(data_path, "igt1_96_withtotalvi20260206_clean_ADTonly.Rds"))@meta.data
seurat_meta_filtered <- seurat_meta[rownames(L_pm_filtered), ]

source("code/R/activation_shared_setup.R")
(3c) Activation effect, CD4 vs. CD8
# ============================================================
# 3a: Standardized mean difference, activated vs resting, CD4 vs CD8
# ============================================================
d_thr <- 0.15
ratio_cutoff <- 3

GP_activation_summary <- diff_factors_merged %>%
  dplyr::inner_join(d_factors_merged %>% dplyr::select(SYMBOL, d_CD4, d_CD8), by = "SYMBOL") %>%
  dplyr::mutate(Ratio_CD8_CD4 = mean_change_loadings_CD8 / mean_change_loadings_CD4) %>%
  dplyr::select(GP = SYMBOL, mean_change_loadings_CD4, mean_change_loadings_CD8, AveExpr_CD4, AveExpr_CD8, d_CD4, d_CD8, Ratio_CD8_CD4)

# Colour every GP using the same four-category rule (ratio + sign + magnitude
# gate via d_thr). Curated GPs (GPs_of_interest) override with their fixed
# manual highlight_colors; non-curated GPs are classified automatically.
# Only the curated GPs are labelled, to keep the plot readable.
manual_curated_df <- GP_activation_summary %>%
  dplyr::mutate(
    auto_color = dplyr::case_when(
      abs(Ratio_CD8_CD4) > ratio_cutoff & abs(d_CD8) > d_thr ~ "darkorange2",
      abs(Ratio_CD8_CD4) < 1 / ratio_cutoff & abs(d_CD4) > d_thr ~ "blue",
      abs(Ratio_CD8_CD4) > 1 / ratio_cutoff & abs(Ratio_CD8_CD4) < ratio_cutoff & d_CD4 > d_thr & d_CD8 > d_thr ~ "darkred",
      abs(Ratio_CD8_CD4) > 1 / ratio_cutoff & abs(Ratio_CD8_CD4) < ratio_cutoff & d_CD4 < -d_thr & d_CD8 < -d_thr ~ "darkgreen",
      TRUE ~ "black"
    ),
    point_color = ifelse(GP %in% GPs_of_interest, highlight_colors[GP], auto_color)
  )

p_3a <- ggplot(manual_curated_df, aes(x = d_CD4, y = d_CD8)) +
  geom_abline(slope = 1, intercept = 0, linetype = "dashed", color = "red") +
  geom_hline(yintercept = 0, linetype = "dashed", color = "blue") +
  geom_vline(xintercept = 0, linetype = "dashed", color = "blue") +
  geom_point(aes(color = point_color), size = 2) +
  ggrepel::geom_text_repel(
    data = filter(manual_curated_df, GP %in% GPs_of_interest),
    aes(label = GP, color = point_color),
    max.overlaps = Inf, size = 3.5, box.padding = 0.35, point.padding = 0.5, segment.color = "grey50"
  ) +
  scale_color_identity() +
  # Signed (pseudo-)log axes: d is signed, so a plain log drops negatives/zeros.
  scale_x_continuous(trans = scales::pseudo_log_trans(sigma = 0.15), breaks = c(-1, -0.5, -0.2, -0.1, 0, 0.1, 0.2, 0.5, 1)) +
  scale_y_continuous(trans = scales::pseudo_log_trans(sigma = 0.15), breaks = c(-1, -0.5, -0.2, -0.1, 0, 0.1, 0.2, 0.5, 1)) +
  coord_equal(xlim = c(-1.6, 1.6), ylim = c(-1.6, 1.6)) +
  labs(
    x = "Standardized Mean Difference (d) for CD4 Activated vs Resting",
    y = "Standardized Mean Difference (d) for CD8 Activated vs Resting",
    title = "GPs colored by semantic category (curated set labeled)"
  ) +
  theme_minimal()
ggsave(filename = paste0(figure_path, "3c.pdf"), plot = p_3a, width = 8, height = 7)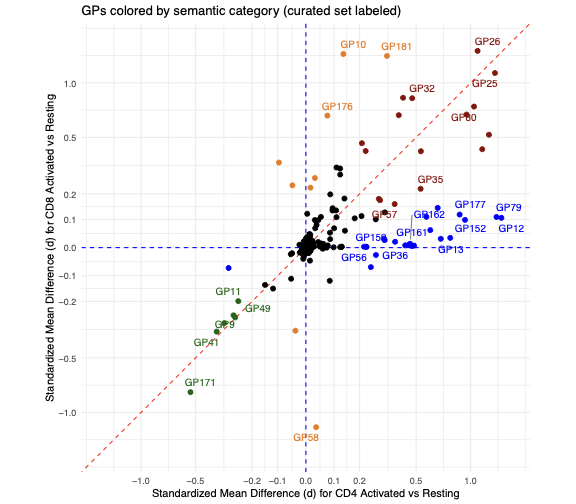
| Version | Author | Date |
|---|---|---|
| 06b2461 | Ziang Zhang | 2026-07-02 |
Fig. 3c. Standardized mean difference (d) in GP loading between activated and resting cells, comparing CD4 (x-axis) and CD8 (y-axis); for each GP, d is the activated-minus-resting difference in mean loading divided by that GP’s loading SD pooled over all activated and resting CD4/CD8 cells. Points are colored based on whether they are highly up-regulated in both CD4/CD8 activation, down-regulated in both, or only in CD4 or CD8 activation; GPs not meeting these criteria are shown in black, and only the curated set is labeled.
(3d) GP-gene signature network
# ============================================================
# 3b: GP-gene signature network
# ============================================================
set.seed(42)
F_pm_filtered_norm_subset <- F_pm_filtered_norm[, GPs_of_interest, drop = FALSE]
top_5_pos <- apply(F_pm_filtered_norm_subset, 2, function(x) {
  idx <- order(abs(x), decreasing = TRUE)[1:5]
  idx <- idx[x[idx] > 0]
  names(x)[idx]
})
top_5_neg <- apply(F_pm_filtered_norm_subset, 2, function(x) {
  idx <- order(abs(x), decreasing = TRUE)[1:5]
  idx <- idx[x[idx] < 0]
  names(x)[idx]
})
names(top_5_pos) <- GPs_of_interest
names(top_5_neg) <- GPs_of_interest

pos_edges <- stack(top_5_pos) %>% dplyr::rename(Gene = values, GP = ind) %>% dplyr::mutate(Type = "Positive", Color = "red")
neg_edges <- stack(top_5_neg) %>% dplyr::rename(Gene = values, GP = ind) %>% dplyr::mutate(Type = "Negative", Color = "blue")
all_edges <- dplyr::bind_rows(pos_edges, neg_edges) %>% dplyr::filter(Gene != "" & !is.na(Gene))
all_edges_sorted <- all_edges %>% dplyr::arrange(Type, GP, Gene)

gp_group_df <- data.frame(
  name = names(highlight_colors),
  ManualGroup = dplyr::case_when(
    highlight_colors == "blue" ~ "CD4 only",
    highlight_colors == "darkorange2" ~ "CD8 only",
    highlight_colors == "darkgreen" ~ "both down",
    highlight_colors == "darkred" ~ "both up",
  )
)
manual_colors_palette <- c("CD4 only" = "blue", "CD8 only" = "darkorange2", "both down" = "darkgreen", "both up" = "darkred", "Gene" = "#666666")

graph <- tidygraph::as_tbl_graph(all_edges_sorted) %>%
  tidygraph::activate(nodes) %>%
  dplyr::mutate(
    NodeGroup = ifelse(name %in% all_edges$GP, "GP", "Gene"),
    Importance = tidygraph::centrality_degree()
  ) %>%
  dplyr::left_join(gp_group_df, by = "name") %>%
  dplyr::mutate(
    ColorGroup = ifelse(NodeGroup == "Gene", "Gene", ManualGroup),
    gp_label = ifelse(NodeGroup == "GP", name, "")
  )

set.seed(2)
p_3b <- ggraph(graph, layout = "stress") +
  geom_edge_link(aes(color = Color), alpha = 0.4, width = 0.6) +
  geom_node_point(aes(filter = (NodeGroup == "Gene"), color = ColorGroup), shape = 16, size = 2, alpha = 0.8) +
  geom_node_point(aes(filter = (NodeGroup == "GP"), color = ColorGroup), shape = 15, size = 10, alpha = 0.7) +
  geom_node_text(aes(filter = (NodeGroup == "GP"), label = gp_label), color = "white", fontface = "bold", size = 3) +
  geom_node_text(aes(filter = (NodeGroup == "Gene"), label = name), repel = TRUE, size = 2.5, color = "black", max.overlaps = 20) +
  scale_edge_color_identity() +
  scale_color_manual(name = "GP Types", values = manual_colors_palette, breaks = c("CD4 only", "CD8 only", "both down", "both up")) +
  theme_void() +
  labs(title = "GP-Gene Signature Network", subtitle = "Nodes colored by manual GP classification", caption = "Red edges: Positive | Blue edges: Negative") +
  theme(legend.position = "bottom", legend.title = element_text(face = "bold"), plot.margin = margin(10, 10, 10, 10)) +
  guides(color = guide_legend(override.aes = list(size = 5, shape = 15)))
ggsave(filename = paste0(figure_path, "3d.pdf"), plot = p_3b, width = 10, height = 10)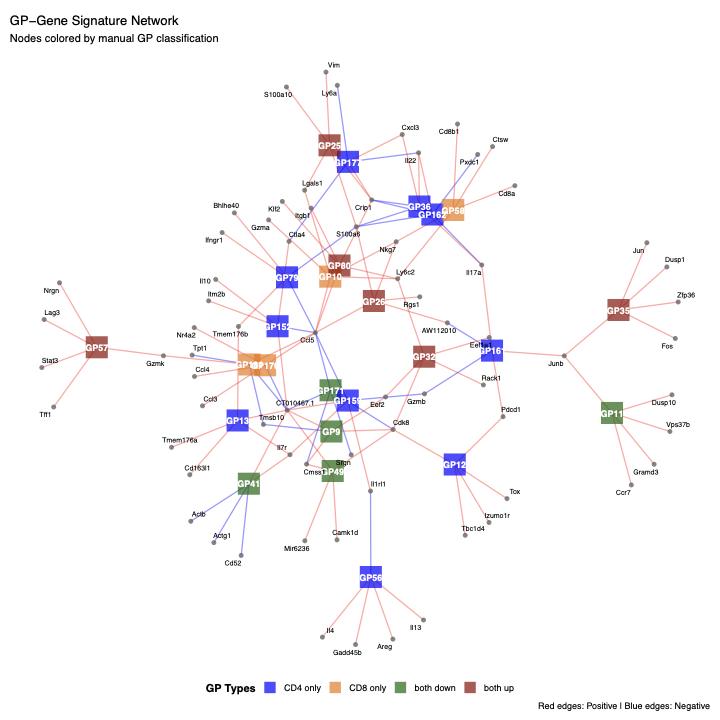
| Version | Author | Date |
|---|---|---|
| 06b2461 | Ziang Zhang | 2026-07-02 |
Fig. 3d. GP-gene signature network in which each GP is linked to its 5 most regulated marker genes (per-GP-normalized gene scores); edges are colored by the score sign (red, positive; blue, negative).
(3e) TF-GP network
# ============================================================
# 3c: Bipartite TF-GP network
# ============================================================
mm <- org.Mm.eg.db::org.Mm.eg.db
go2eg <- as.list(org.Mm.eg.db::org.Mm.egGO2ALLEGS)
tf_symbols <- AnnotationDbi::select(mm, keys = unique(unlist(go2eg)), columns = "SYMBOL", keytype = "ENTREZID")
tf <- c(
  sort(tf_symbols$SYMBOL[tf_symbols$ENTREZID %in% unique(go2eg[["GO:0003700"]])]),
  "Tox", "Tox2", "Tox3", "Tox4"
) %>% sort() %>% unique()

F_sub_tf <- F_pm_filtered_norm[, GPs_of_interest, drop = FALSE]
tf_gp_threshold <- 0.25
tf_in_F <- intersect(tf, rownames(F_sub_tf))
tf_max_score <- apply(F_sub_tf[tf_in_F, , drop = FALSE], 1, max, na.rm = TRUE)
selected_tfs <- sort(names(tf_max_score)[tf_max_score > tf_gp_threshold])

# Top-to-bottom group order in the network: both-up -> CD8-only -> both-down -> CD4-only
gp_color_group_order <- c("darkred", "darkorange2", "darkgreen", "blue")

tf_network_plot <- plot_tf_gp_network_v2(
  F = F_sub_tf,
  selected_tfs = selected_tfs,
  tf_gp_threshold = tf_gp_threshold,
  top_genes_per_gp = 5,
  gp_colors = highlight_colors,
  gp_group_order = gp_color_group_order,
  optimize_layout = TRUE,
  barycenter_iter = 12,
  gp_spacing = 1.5,
  node_size_tf = 6,
  node_size_gp = 5,
  label_size_tf = 4.5,
  label_size_gp = 4,
  label_size_gene = 3.4
)
plot_height_tf <- min(60, max(12, length(selected_tfs) * 0.35, length(GPs_of_interest) * 1.5 * 0.55 + 2))
ggsave(filename = paste0(figure_path, "3e.pdf"), plot = tf_network_plot, width = 18, height = plot_height_tf, limitsize = FALSE)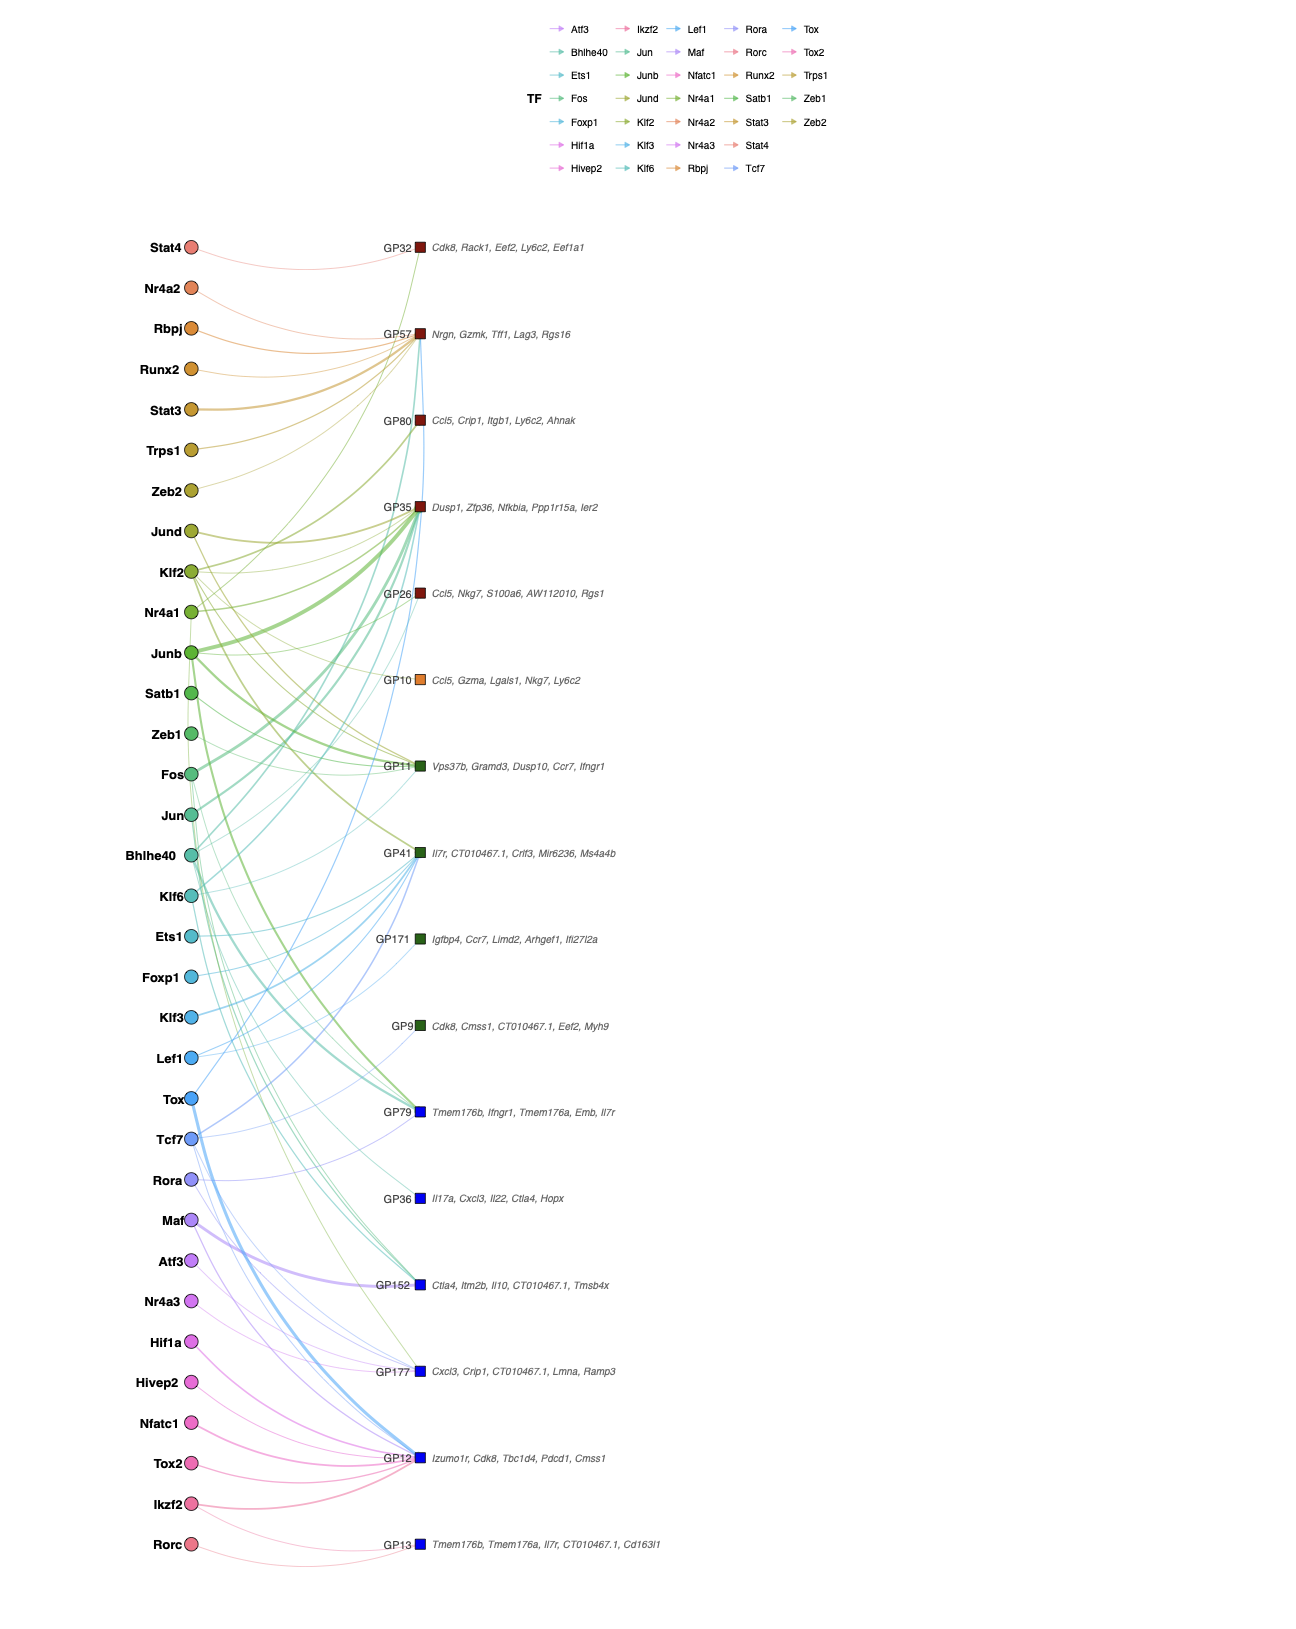
| Version | Author | Date |
|---|---|---|
| 06b2461 | Ziang Zhang | 2026-07-02 |
Fig. 3e. Bipartite transcription factor (TF)-GP network: TFs (GO:0003700 DNA-binding TFs plus the Tox family) with a normalized score exceeding 0.25 in any curated GP are connected to those GPs by directed edges (width proportional to the normalized score, color identifying the TF); the top 5 positively regulated genes per GP are listed beside each GP node.
(3f) Log2FC heatmap across conditions
# ============================================================
# 3d: log2FC heatmap of activated CD4+CD8 cells across conditions,
#     relative to the per-GP mean across all CD4/CD8 cells
# ============================================================
act_keep <- seurat_meta_filtered$annotation_level1 %in% c("CD4", "CD8") & seurat_meta_filtered$annotation_level2_group == "activated"
meta_act <- seurat_meta_filtered[act_keep, ]
L_act <- L_subset[act_keep, , drop = FALSE]

min_cells_cond <- 50
cd_lin <- table(meta_act$condition_detailed_simplified, meta_act$annotation_level1)
cond_keep <- rownames(cd_lin)[cd_lin[, "CD4"] >= min_cells_cond & cd_lin[, "CD8"] >= min_cells_cond]

cd_br <- table(meta_act$condition_detailed_simplified, meta_act$condition_broad)
cd_to_broad <- setNames(colnames(cd_br)[apply(cd_br, 1, which.max)], rownames(cd_br))[cond_keep]

# Column order: `healthy` broad first (with `baseline` as its first condition)
broad_rank <- ifelse(cd_to_broad == "healthy", 0L, 1L)
within_broad_rank <- ifelse(cd_to_broad == "healthy" & cond_keep == "baseline", 0L, 1L)
col_order_cond <- order(broad_rank, cd_to_broad, within_broad_rank, cond_keep)
cond_keep <- cond_keep[col_order_cond]
cd_to_broad <- cd_to_broad[cond_keep]

mean_mat_cond <- vapply(cond_keep, function(cond) colMeans(L_act[meta_act$condition_detailed_simplified == cond, , drop = FALSE]), numeric(ncol(L_act)))
mean_mat_cond <- mean_mat_cond[gp_row_order, , drop = FALSE]
broad_levels <- unique(cd_to_broad)
col_anno_cond <- data.frame(condition_broad = factor(cd_to_broad, levels = broad_levels), row.names = colnames(mean_mat_cond))
row_label_cols <- group_colors[gp_to_group[gp_row_order]]

pc_lfc <- 1e-10
cap_lfc <- 2
cd4cd8_idx <- seurat_meta_filtered$annotation_level1 %in% c("CD4", "CD8")
L_cd4cd8 <- L_subset[cd4cd8_idx, , drop = FALSE]
mu_lfc_mean <- colMeans(L_cd4cd8, na.rm = TRUE)[rownames(mean_mat_cond)]
lfc_mat_mean <- log2((mean_mat_cond + pc_lfc) / (mu_lfc_mean + pc_lfc))
lfc_mat_mean <- pmax(pmin(lfc_mat_mean, cap_lfc), -cap_lfc)

ph_cond_lfc_mean <- pheatmap(
  lfc_mat_mean, cluster_rows = FALSE, cluster_cols = FALSE,
  color = colorRampPalette(c("#7A0177", "black", "#FFD700"))(101),
  breaks = seq(-cap_lfc, cap_lfc, length.out = 102),
  annotation_col = col_anno_cond,
  gaps_row = head(cumsum(lengths(gp_groups)), -1),
  gaps_col = head(cumsum(rle(as.character(cd_to_broad))$lengths), -1),
  main = "log2FC vs per-GP MEAN across all CD4/CD8 (activated CD4+CD8 by condition_detailed_simplified)",
  silent = TRUE
)
row_idx_lfc_m <- which(ph_cond_lfc_mean$gtable$layout$name == "row_names")
ph_cond_lfc_mean$gtable$grobs[[row_idx_lfc_m]]$gp$col <- row_label_cols

pdf(paste0(figure_path, "3f.pdf"), width = max(8, 0.18 * ncol(lfc_mat_mean) + 4), height = 6)
grid::grid.draw(ph_cond_lfc_mean$gtable)
invisible(dev.off())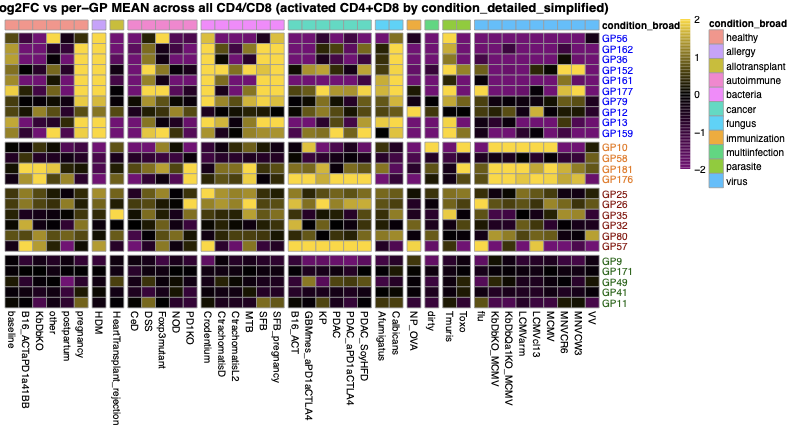
| Version | Author | Date |
|---|---|---|
| 06b2461 | Ziang Zhang | 2026-07-02 |
Fig. 3f. Heatmap of log2 fold-change in mean GP loading across experimental conditions for activated CD4 and CD8 cells, computed relative to each GP’s mean loading across all CD4/CD8 cells (resting and activated, all conditions) and capped at +-2, with a small pseudocount (1e-10) added to stabilize computation; columns are conditions with >=50 activated CD4 and >=50 activated CD8 cells, grouped and annotated by broad condition category, with color from purple (low) through black (no change) to gold (high).
(3g) Mean loading by sub-lineage
# ============================================================
# 3e: Mean GP loading per Level-2 sub-lineage (built before 3d since 3d
#     reuses gp_row_order/group_colors computed here)
# ============================================================
keep_cells <- seurat_meta_filtered$annotation_level1 %in% lineages
meta_sub <- seurat_meta_filtered[keep_cells, ]
L_keep <- L_subset[keep_cells, , drop = FALSE]

l2_counts <- table(meta_sub$annotation_level2)
l2_keep <- names(l2_counts)[l2_counts >= 50]
# Drop the "P" cluster and any "w..." clusters (wM, wW, etc.) across all lineages
l2_stripped <- sub("^[^._]+[._]", "", l2_keep)
exclude_l2 <- l2_stripped == "P" | grepl("^w", l2_stripped, ignore.case = TRUE) | grepl("[._]w", l2_keep, ignore.case = TRUE)
l2_keep <- l2_keep[!exclude_l2]

mean_mat <- vapply(l2_keep, function(l2) colMeans(L_keep[meta_sub$annotation_level2 == l2, , drop = FALSE]), numeric(ncol(L_keep)))
l2_to_l1 <- vapply(l2_keep, function(l2) as.character(meta_sub$annotation_level1[meta_sub$annotation_level2 == l2][1]), character(1))
col_order <- order(match(l2_to_l1, lineages), l2_keep)
mean_mat <- mean_mat[gp_row_order, col_order]
l2_to_l1 <- l2_to_l1[col_order]

immgen_cols <- ZemmourLib::immgent_colors
col_anno <- data.frame(Lineage = factor(l2_to_l1, levels = lineages), row.names = colnames(mean_mat))
anno_colors_mean <- list(Lineage = immgen_cols$level1[lineages])
row_label_cols <- group_colors[gp_to_group[gp_row_order]]
col_label_cols <- immgen_cols$level2[colnames(mean_mat)]
col_label_cols[is.na(col_label_cols)] <- "black"

ph <- pheatmap(
  mean_mat, cluster_rows = FALSE, cluster_cols = FALSE,
  color = colorRampPalette(c("white", "red"))(200),
  annotation_col = col_anno, annotation_colors = anno_colors_mean,
  gaps_row = head(cumsum(lengths(gp_groups)), -1),
  gaps_col = head(cumsum(rle(l2_to_l1)$lengths), -1),
  main = "Average loading of Figure 3 GPs per Level-2 sub-lineage",
  silent = TRUE
)
row_idx <- which(ph$gtable$layout$name == "row_names")
col_idx <- which(ph$gtable$layout$name == "col_names")
ph$gtable$grobs[[row_idx]]$gp$col <- row_label_cols
ph$gtable$grobs[[col_idx]]$gp$col <- col_label_cols

pdf(paste0(figure_path, "3g.pdf"), width = 11, height = 5.5)
grid::grid.draw(ph$gtable)
invisible(dev.off())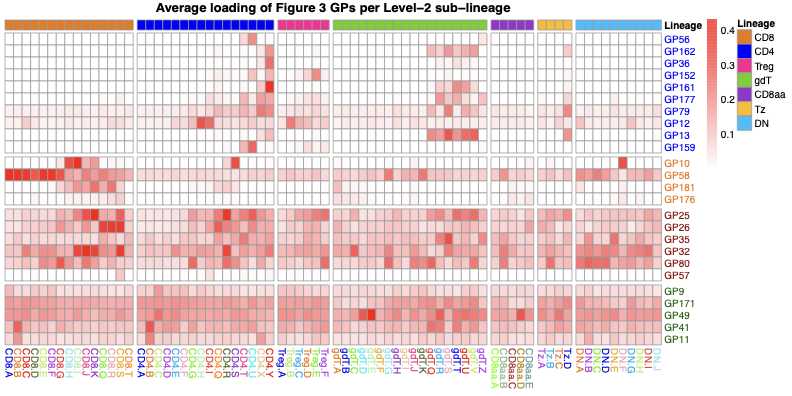
| Version | Author | Date |
|---|---|---|
| 06b2461 | Ziang Zhang | 2026-07-02 |
Fig. 3g. Heatmap of mean GP loading per Level-2 sub-lineage across the seven T-cell lineages (CD8, CD4, Treg, gdT, CD8aa, Tz, DN; sub-types with >=50 cells), with the top bar denoting parent lineage and color from white (low) to red (high).
sessionInfo()R version 4.5.1 (2025-06-13)
Platform: aarch64-apple-darwin20
Running under: macOS Sequoia 15.6.1
Matrix products: default
BLAS: /System/Library/Frameworks/Accelerate.framework/Versions/A/Frameworks/vecLib.framework/Versions/A/libBLAS.dylib
LAPACK: /Library/Frameworks/R.framework/Versions/4.5-arm64/Resources/lib/libRlapack.dylib; LAPACK version 3.12.1
locale:
[1] en_CA/en_CA/en_CA/C/en_CA/en_CA
time zone: Asia/Tokyo
tzcode source: internal
attached base packages:
[1] stats graphics grDevices utils datasets methods base
loaded via a namespace (and not attached):
[1] vctrs_0.7.3 cli_3.6.6 knitr_1.50 rlang_1.2.0
[5] xfun_0.55 stringi_1.8.7 otel_0.2.0 promises_1.5.0
[9] jsonlite_2.0.0 workflowr_1.7.2 glue_1.8.1 rprojroot_2.1.1
[13] git2r_0.36.2 htmltools_0.5.9 httpuv_1.6.16 sass_0.4.10
[17] rmarkdown_2.30 evaluate_1.0.5 jquerylib_0.1.4 tibble_3.3.0
[21] fastmap_1.2.0 yaml_2.3.12 lifecycle_1.0.5 whisker_0.4.1
[25] stringr_1.6.0 compiler_4.5.1 fs_1.6.6 Rcpp_1.1.1-1.1
[29] pkgconfig_2.0.3 later_1.4.4 digest_0.6.39 R6_2.6.1
[33] pillar_1.11.1 magrittr_2.0.5 bslib_0.9.0 tools_4.5.1
[37] cachem_1.1.0